Guidance of CCMS
Greetings

Supercomputing System for Computational Materials Science
Mission of Center for Computational Materials Science
System Overview
System Configuration
MASAMUNE-II consists of the Supercomputer (HPE Cray XD220v, HPE Scale-up Server 3200, and HPE Cray XD670). It is integrated by Hitachi, Ltd. The system specifications are shown below.
| System Name | Supercomputer | |||
|---|---|---|---|---|
| Model | HPE Cray XD220v | HPE Scale-up Server 3200 | HPE Cray XD670 | |
| total nodes | General node:120 |
2 nodes |
11 nodes |
2 nodes |
| CPU |
|
Intel Xeon Platinum 8490H
|
Intel Xeon Platinum 8480+
|
Intel Xeon Platinum 8570+
|
| Accelerator | - | - |
NVIDIA H100 80GB SXM5
|
NVIDIA H200 141GB SXM5
|
| memory size | 512 GiB/node 2.0 TiB/node |
4.0 TiB/node | 1.0 TiB/node | 1.0 TiB/node |
| Theoretical peak performance | 4.603 PFLOPS (CPU : 1.119 PFLOPS, GPU : 3.484 PFLOPS) |
|||
Nickname and Front Panel Design of Supercomputing System
Nickname: MASAMUNE-II
MAterials science Supercomputing system for Advanced MUlti-scale simulations towards NExt-generation - II

Nickname “MASAMUNE-II” has been given to the supercomputing system of CCMS, continuing the nickname of the previous system “MASAMUNE-IMR”, in honor of “Masamune Date”, the first lord of Sendai in the 17th century, who dispatched the sailing ship “San Juan Bautista” towards the world from Sendai in 1613. The nickname “MASAMUNE-II” expresses our hope that creative and innovative achievements of the advanced multi-scale simulations on materials science by our supercomputing system will give great impacts towards the next-generation from Sendai to the World. “Masamune Date” is drawn on the front panel of our supercomputer by Sumi-E Artist OKAZU with the brave letter “弐” meaning “second” in Japanese Kanji. This “Masamune Date” on the front panel is looking ahead to the future advances in the materials science of the world from Sendai.
Achievement
Elucidation of Effect of Environments on Wear Phenomena of Diamond-like Carbon and Proposal of Design Princples towards Improving its Durability
Diamond-like carbon is an amorphous carbon material which has excellent properties such as ultra-low friction, wear resistance, and chemical stability and then it is expected to be used in space stations, airplane engines, drones, medical equipment, etc. However, the wear of diamond-like carbon can lead to reduced material lifespan, machine failure, and even unexpected accidents. Therefore, minimizing its wear to the utmost is a critical challenge for realizing a safe and secure society. Especially, its wear behavior varies signi cantly even with slight environmental differences such as partial pressures of water vapor, oxygen, nitrogen, and hydrogen, and then experimental research has not yet advanced our understanding of its underlying mechanism. Therefore, it is strongly demanded to utilize computational science simulations for clarifying the effects of environments and atmosphere on its wear mechanism and then for establishing the design guidelines to reduce its wear. However, unlike “material-intrinsic properties” such as hardness and electrical conductivity, the changes in wear phenomena due to the environments represent “material response characteristics”.
Therefore, understanding multi-physics phenomena including chemical reactions, friction, impact, stress, fluids, and heat transfer by computational science simulations is essential, which is one of the challenging tasks in computational science simulations. Therefore, by developing a reactive molecular dynamics simulator capable of elucidating complex multi-physics phenomena involving chemical reactions on the supercomputer “MASAMUNE-IMR,” for the first time in the world we discovered that wear occurs via two mechanisms, chemical wear and mechanical wear, on diamond-like carbon and revealed that the chemical reactions occurring on the surface of diamond-like carbon during friction vary depending on the environments such as water and oxygen, leading to differences in the form and amount of chemical wear and mechanical wear. This achievement contributes to extending the lifespan of mechanical systems as well as preventing failures and accidents.
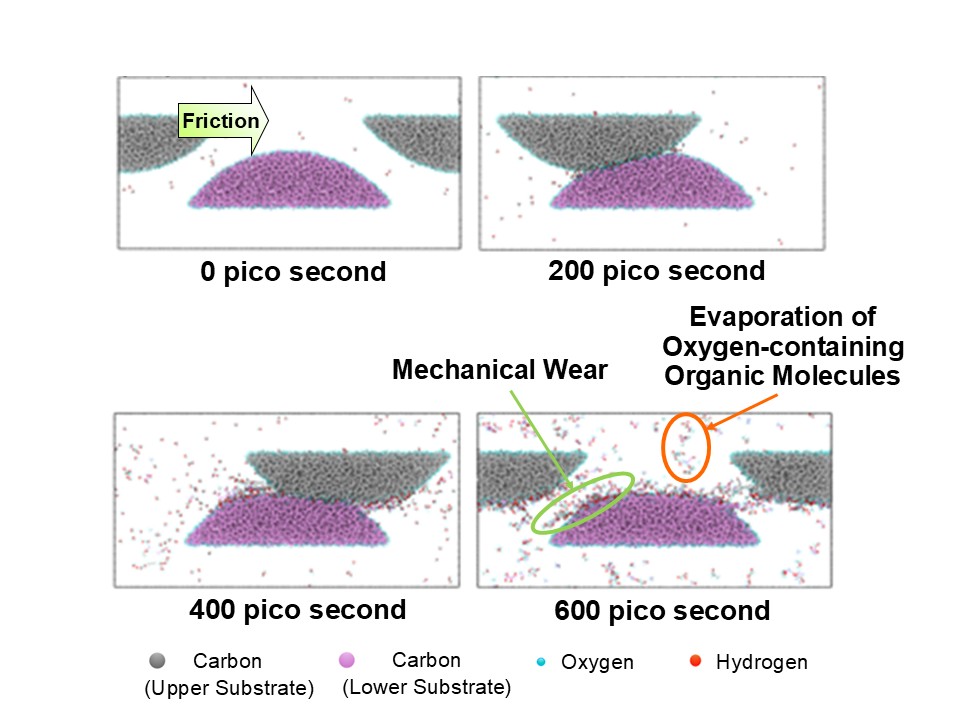
Figure. Friction simulation of diamond-like carbon under water environment (Water hidden for clarity). We have demonstrated that water molecules dissociate into H+ and OH- at carbon atoms with dangling bonds on diamond-like carbon surfaces, initiating chemical wear where oxygen-containing organic molecules such as methanol and ethanol evaporate.
Jing Zhang, Yang Wang, Qian Chen, Yixin Su, Shandan Bai, Yusuke Ootani, Nobuki Ozawa, Koshi Adachi and Momoji Kubo
Carbon, 231 (2025) 119713, DOI: https://doi.org/10.1016/j.carbon.2024.119713
High-Tc Superconductivity in an Effective Hamiltonian for Bilayer Nickelates
High-temperature (high-Tc) superconductivity has recently been reported in the bilayer nickelate c under high pressure. In this study, we focused on the bilayer Hubbard model as a minimal model for its relevant electronic states. The model describes electrons moving on two coupled square-lattice layers with strong repulsion between electrons occupying the same site. Despite its simple form, accurately solving the model is challenging because of strong electron correlations.
We performed large-scale numerical calculations on a supercomputer using cluster dynamical mean-field theory, which accurately treats dynamical electron correlations. The calculations revealed high-Tc superconductivity and showed that strong correlations play an essential role in its emergence.
The metallic state above Tc also exhibits unusual behavior. As shown in the lower panel of Figure, correlation effects become more pronounced from right to left, and part of the low-energy electronic spectral weight is strongly suppressed in the strongly correlated regime. This pseudogap behavior is also a characteristic feature of high-Tc cuprate superconductors, suggesting that nickelates and cuprates may share common physics originating from strong electron correlations.
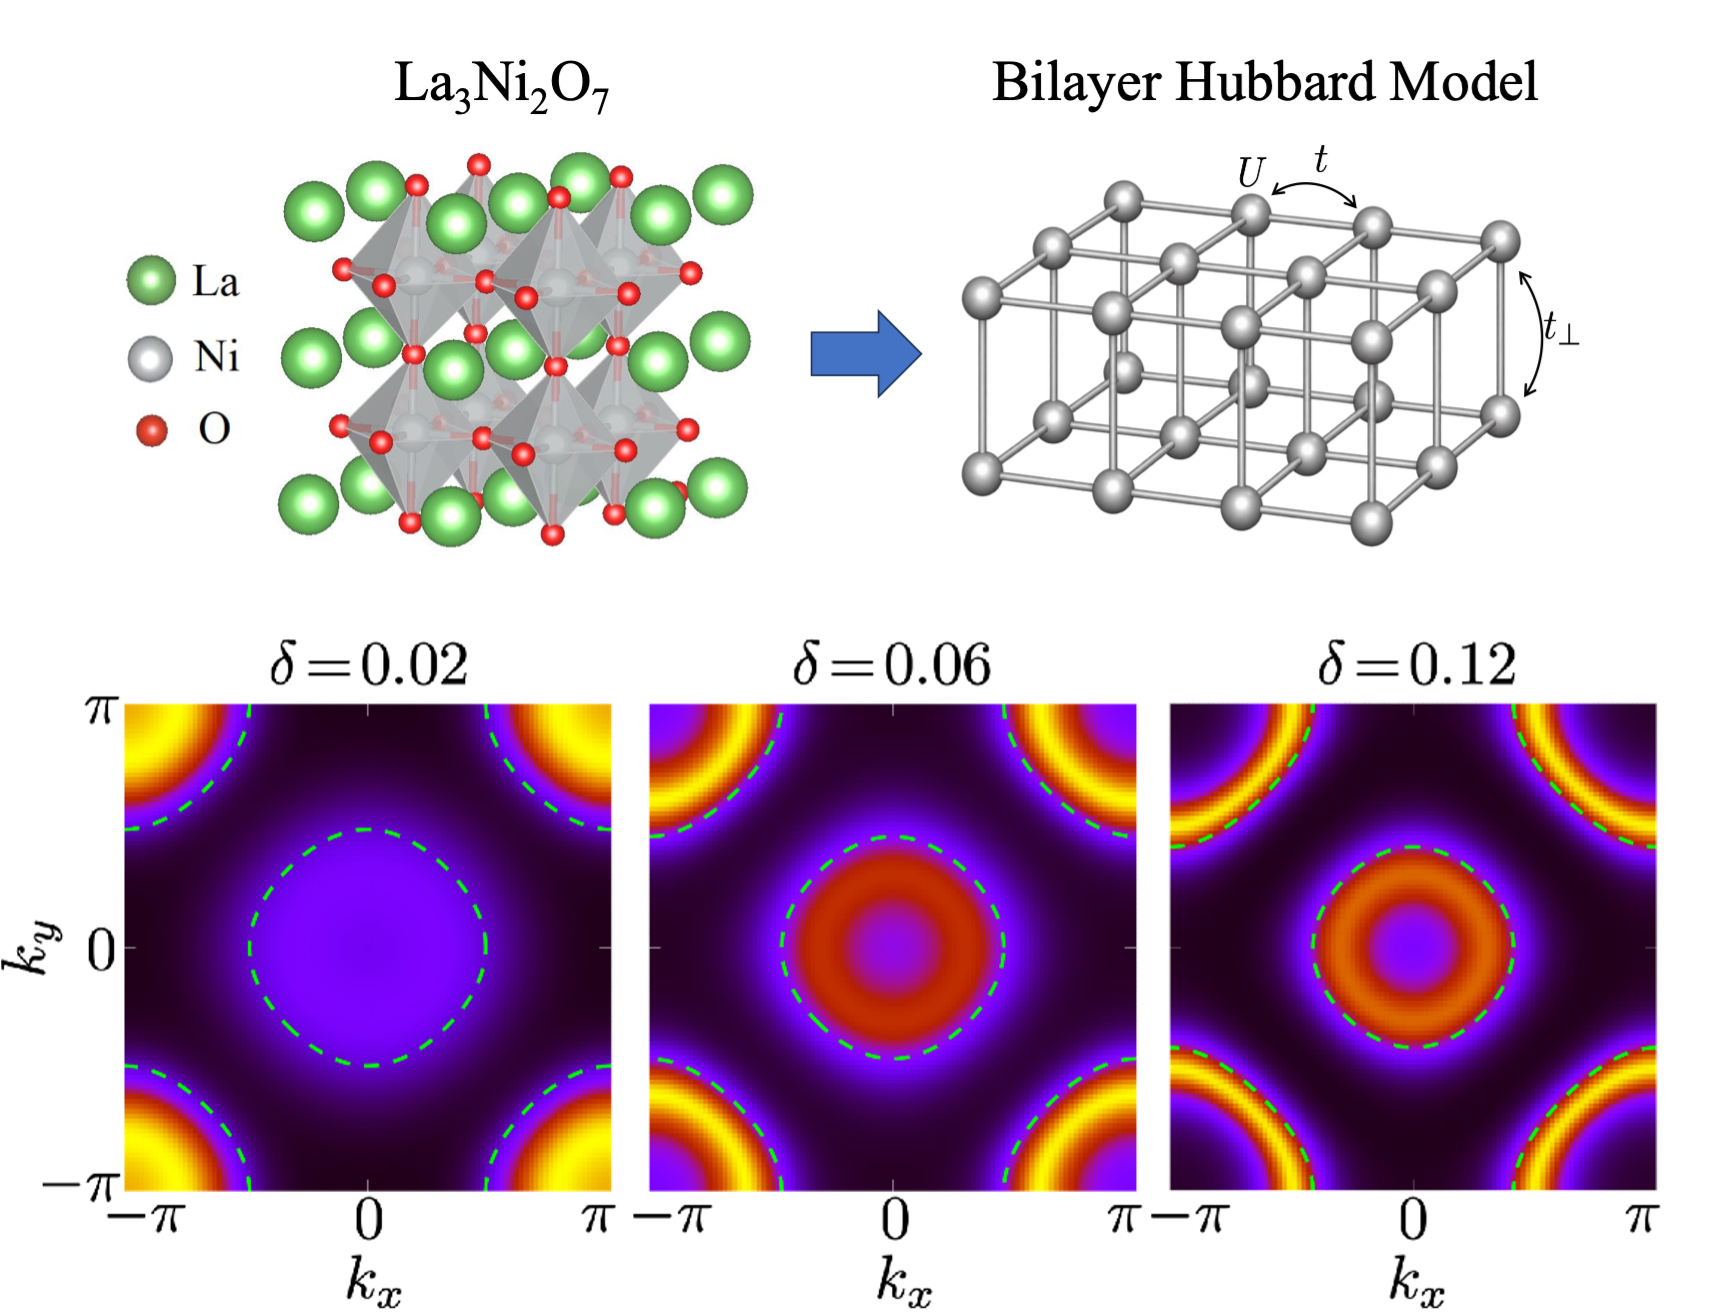
Figure. Bilayer Hubbard model for La3Ni2O7 and its electronic states. The upper panel shows the model derived from the crystal structure, while the lower panel shows the momentum distribution of low-energy electronic states. In the strongly correlated regime (left), a pseudogap appears.
Yusuke Nomura, Motoharu Kitatani, Shiro Sakai and Ryotaro Arita
Phys. Rev. B, 112 (2025) L020504, DOI: https://doi.org/10.1103/dygc-94fq
Balancing Reactivity and Ion Transport in Mg Alloy Anodes via Secondary- Phase Engineering
MSolid-state Mg metal batteries are promising for safe and high-energy storage, yet their performance is limited by unstable anode/electrolyte interfaces. The key challenge is to regulate, rather than eliminate, interfacial reactions: moderate reactions generate Mg2+ transport pathways, whereas excessive reactions cause passivation and polarization. This work shows that secondaryphase engineering can balance interfacial reactivity and transport in Mg alloy anodes. Density functional theory calculations establish a descriptor-guided framework for alloy/electrolyte interface design. Interface formation energies quantify the reaction tendency between Mg-alloy secondary phases and a Mg(BH4)2-based solid electrolyte, while charge density difference analysis reveals electron redistribution and bonding interactions. These calculations identify Mg2Sn as a favorable secondary phase with controlled reactivity and reduced passivation risk. The resulting percolated Mg2Sn network with continuous α-Mg pathways supports interfacial stability and Mg2+ transport. This work connects atomic-scale interface chemistry with alloy anode design and highlights controlled reactivity as a key principle for stable Mg alloy anodes.
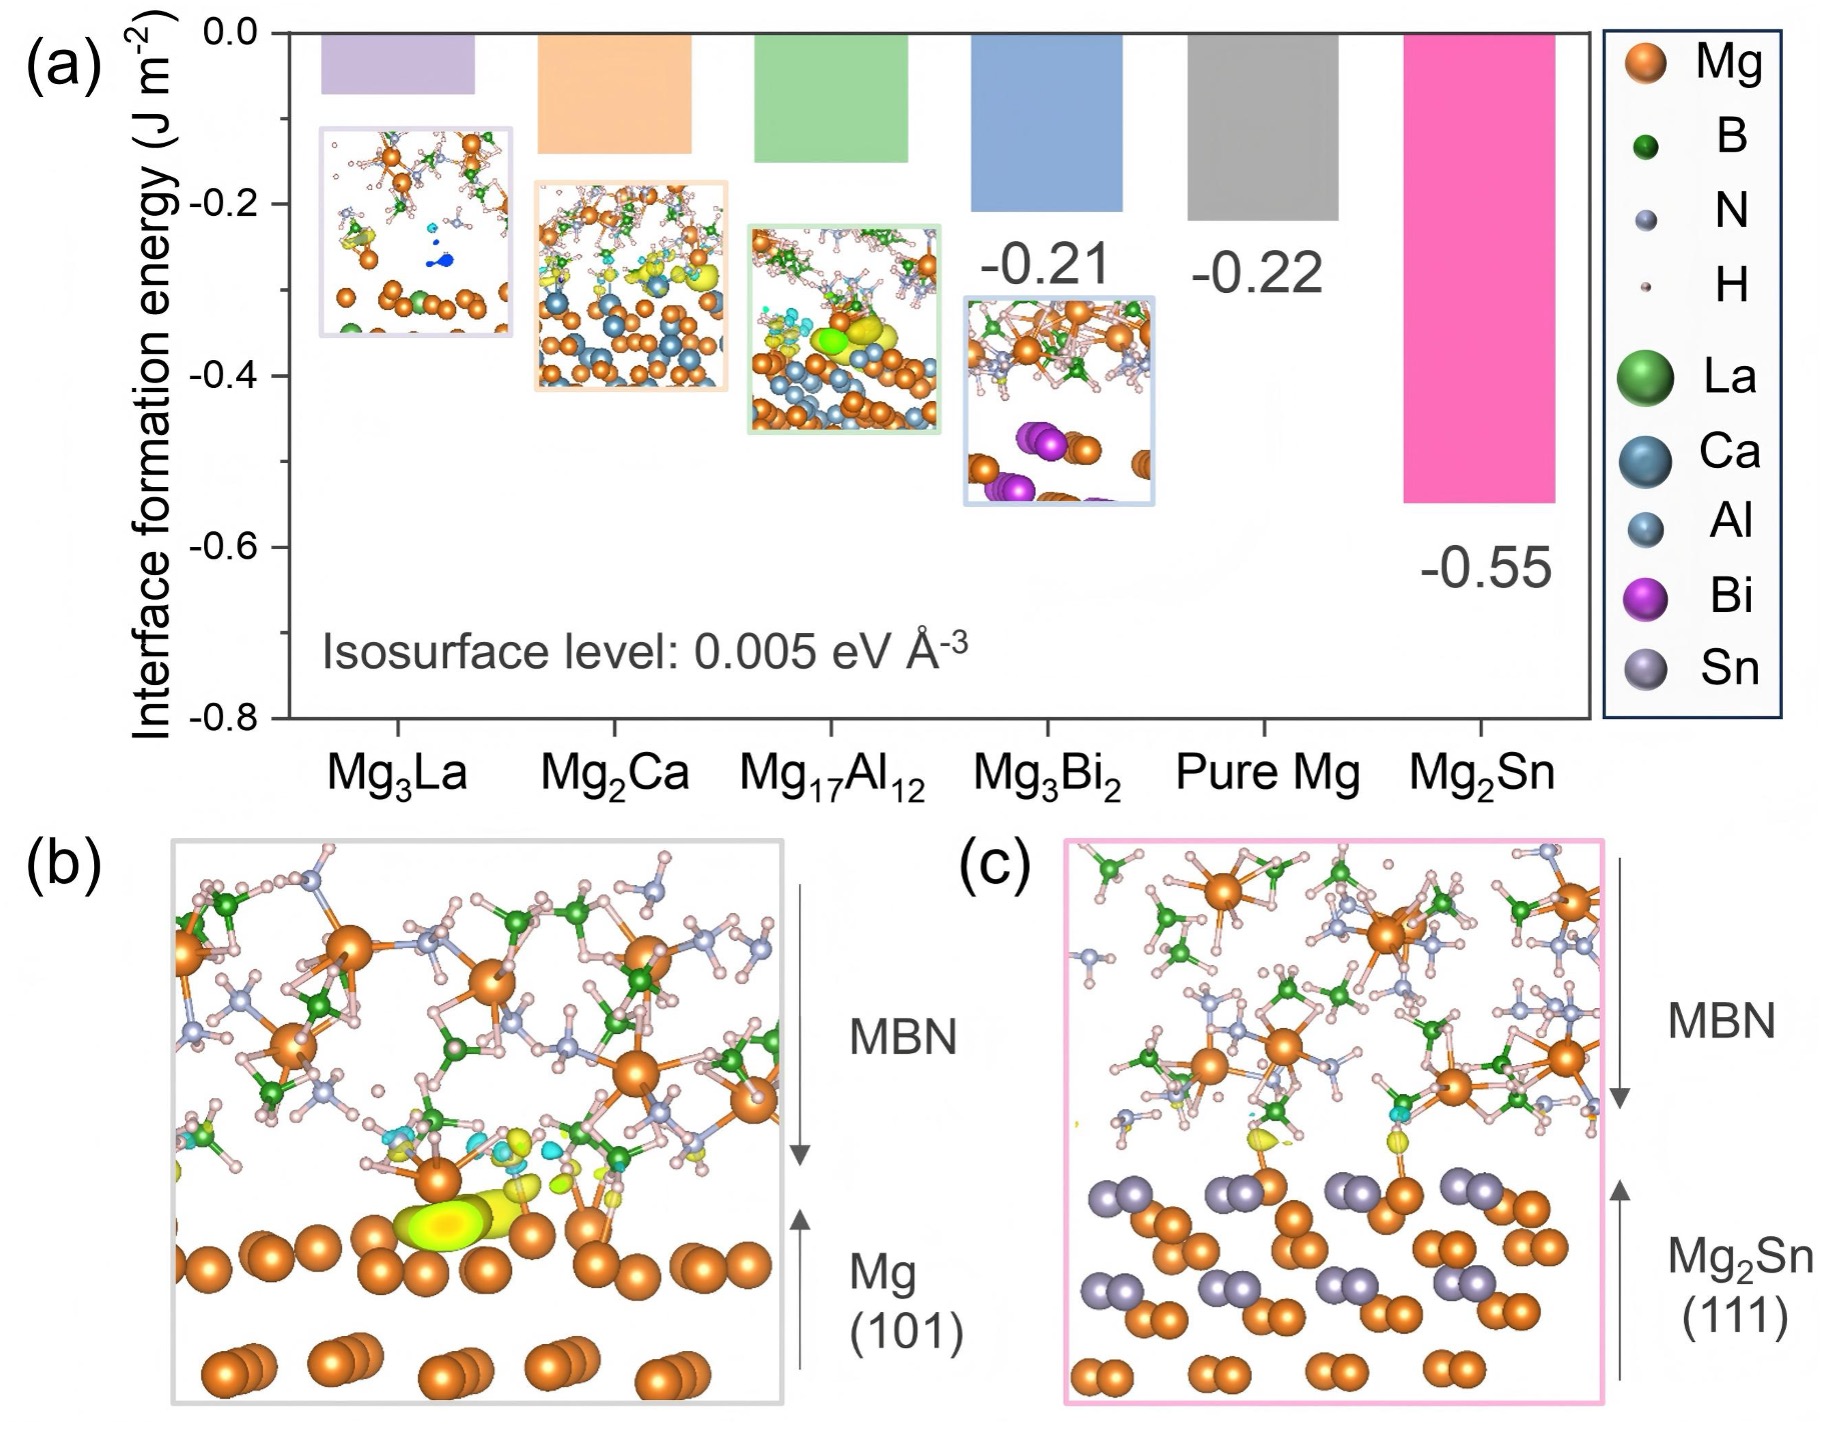
Figure. Reactivity and Mg2+ transport at Mg-alloy/solid-electrolyte interfaces regulated through secondary-phase engineering.
(a) Interface formation energies of various Mg-alloy/MBN interfaces and atomic structures of (b) Mg(101)/MBN and (c) Mg2Sn(111)/MBN interfaces.
MBN: Mg(BH4)2·2NH3.
Qian Wang, Xue Jia, Ting Xu, Jianwei Li, Yungui Chen, Hao Li and Yigang Yan
ACS Energy Lett., 11 (2026) pp.4621-4628, DOI: https://doi.org/10.1021/acsenergylett.6c00909
Lattice Oxygen Insertion Mechanism in Cerium Oxide Catalysts
Cerium oxide (CeO2) is an important catalyst widely used in organic synthesis as well as environmental and energy-related applications. However, the molecular-level origin of its high catalytic activity has remained largely unclear. In this study, we employed first-principles molecular dynamics simulations in conjunction with enhanced sampling techniques to investigate the nitrile hydration reaction in aqueous solution. We discovered a previously unknown reaction mechanism in which a surface lattice oxygen atom, rather than an oxygen atom originating from the solvent water, is incorporated directly into the product. This theoretical prediction was further verified experimentally using oxygen isotope labeling. The direct participation of lattice oxygen has generally been considered to occur only in high-temperature reactions, but our study provides the evidence that lattice oxygen can actively participate in catalytic reactions even under mild conditions near room temperature.
The free energy calculations required extensive first-principles molecular dynamics simulations, which were enabled by the massively parallel computing environment of MASAMUNE-IMR. This computational platform made it possible to elucidate complex solid–liquid interfacial reactions at the atomic level. The findings of this study not only advance our understanding of the reaction mechanism of CeO2 catalysts but also provide valuable insights into the rational design of high-performance catalysts.
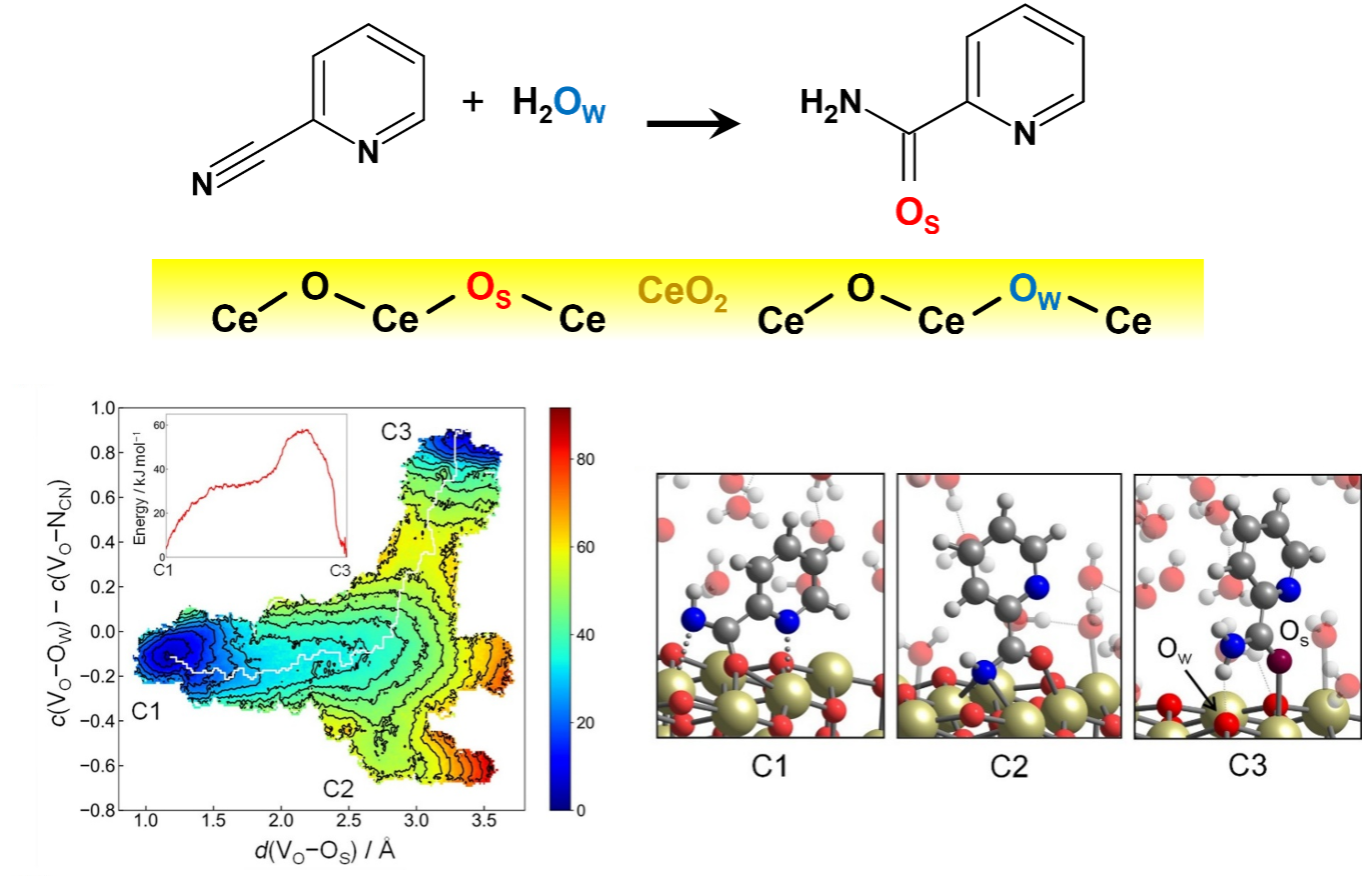
Figure. Free energy analysis of the nitrile hydration reaction over a CeO2 catalyst. First-principles molecular dynamics simulations revealed that the reaction pathway involving incorporation of a surface lattice oxygen atom into the product proceeds with a lower free energy barrier than the conventional pathway involving an oxygen atom derived from water.
Takaaki Endo, Tatsushi Ikeda, Koki Muraoka, Yusuke Kita, Masazumi Tamura and Akira Nakayama
Chem. Sci., 16 (2025) pp.939-951, DOI: https://doi.org/10.1039/d4sc06294a
Access
Access from Sendai Station
walk (20 min)
Taxi (10 min)
Subway (10-15 min)
From Sendai Airport
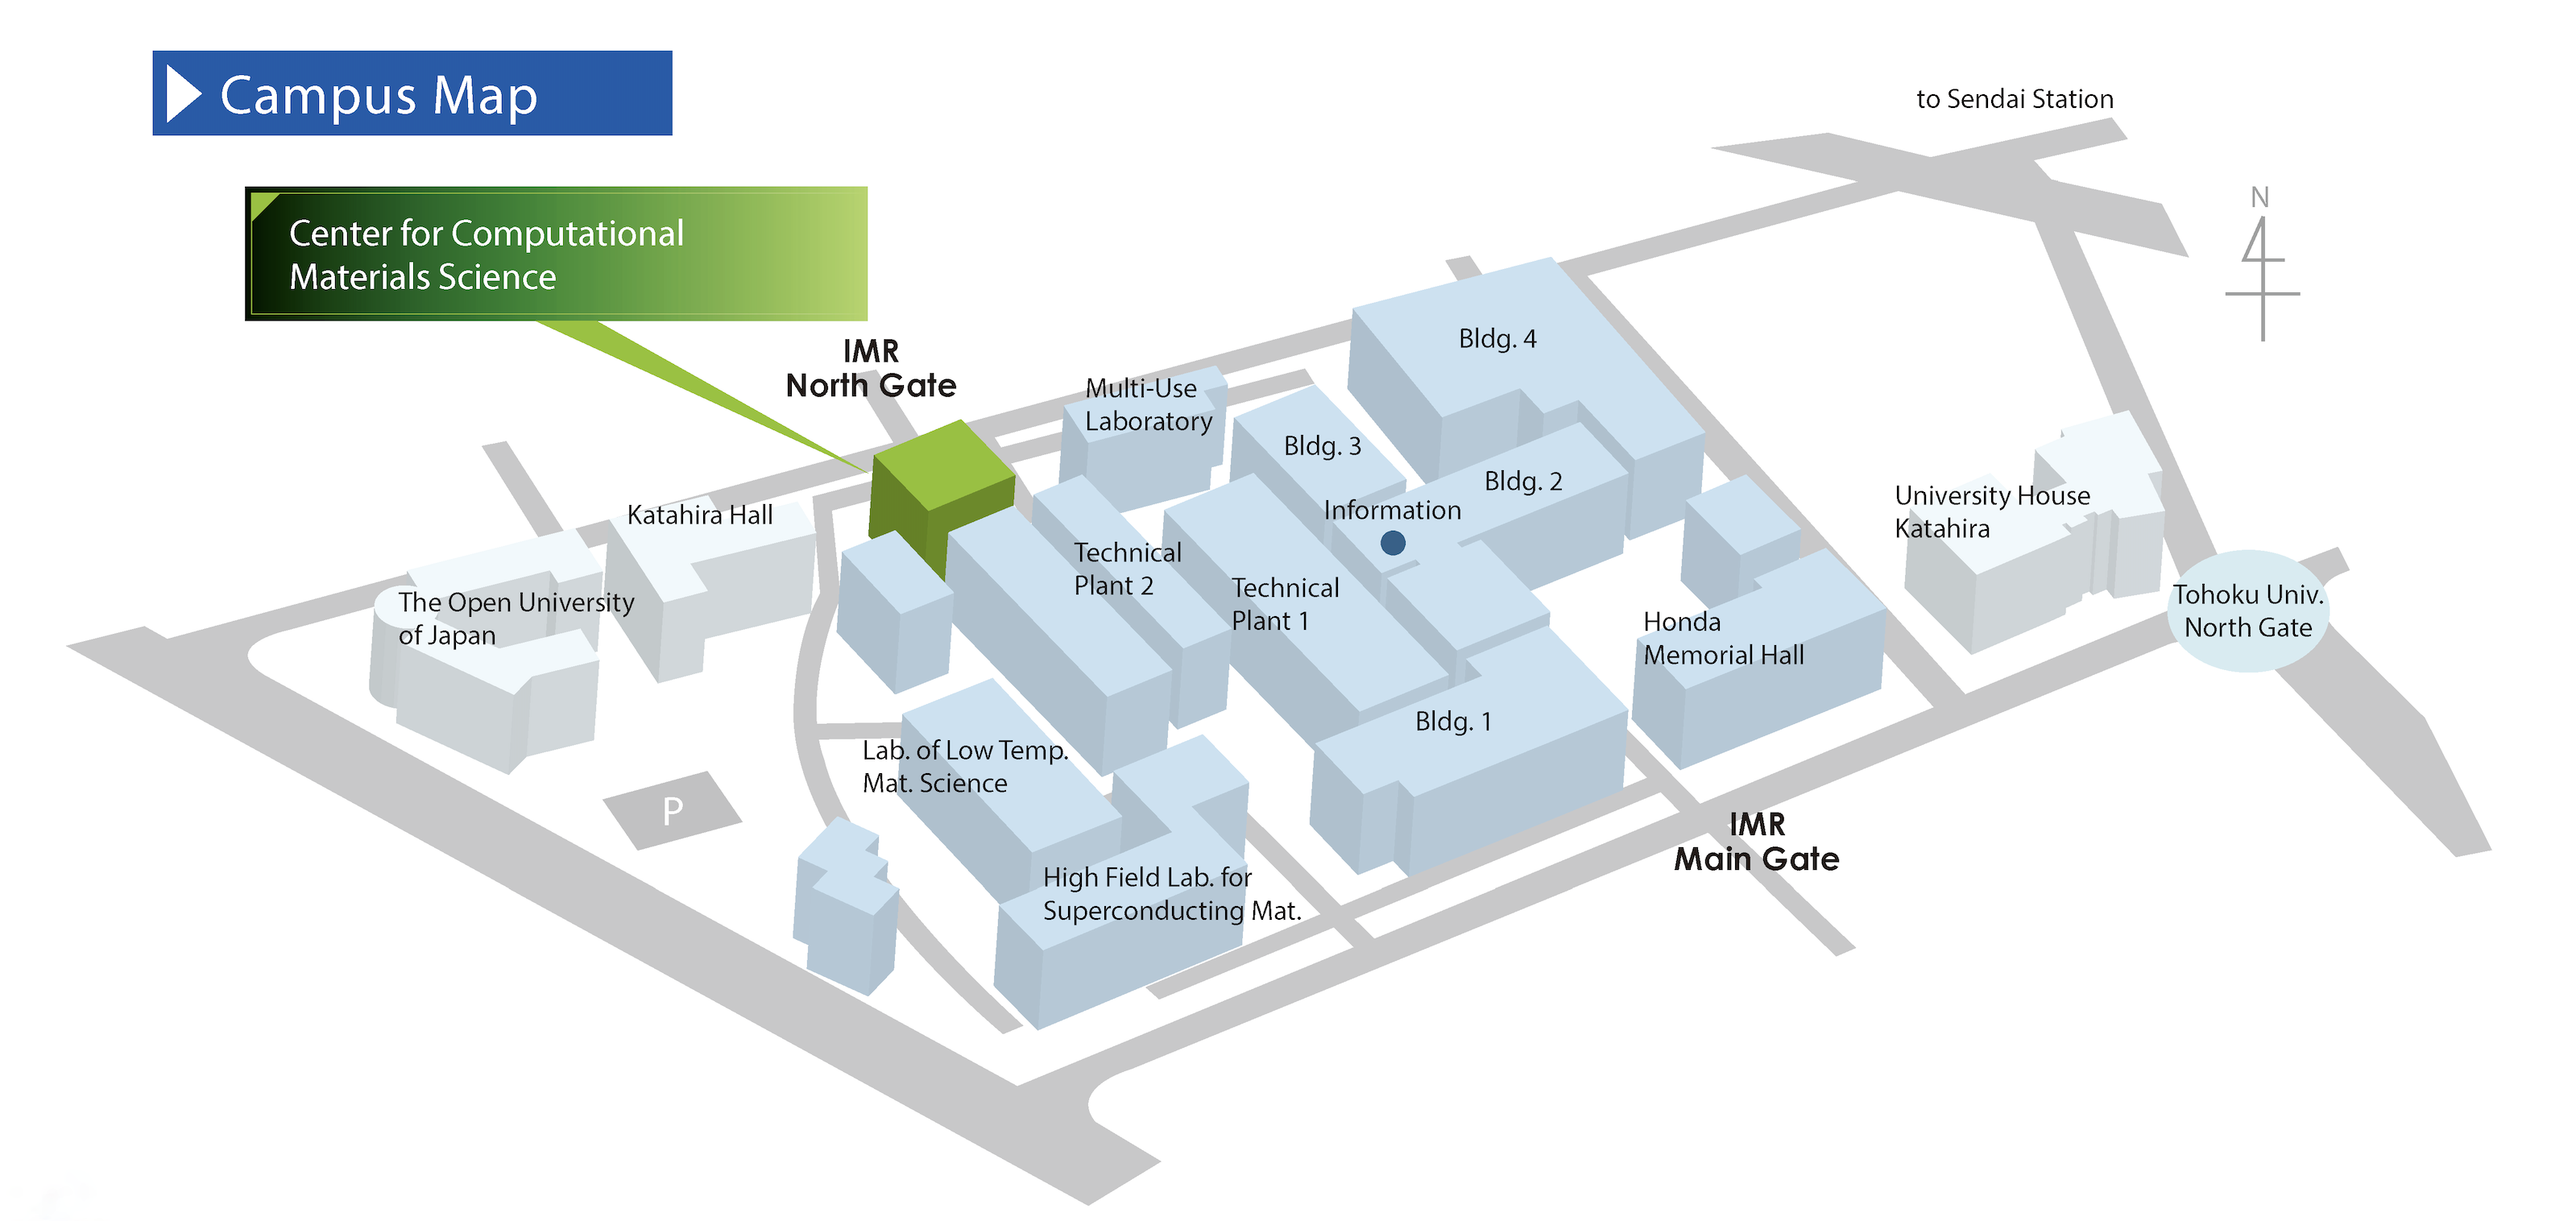
Institute for Materials Research, Tohoku University
Center for Computational Materials Science
TEL: +81-22-215-2411, FAX: +81-22-215-2166